LIVE WEBINAR
Share:

EU MDR Technical Documentation: Keeping Risk, Clinical and Design Records Consistent Through Product Changes

  • Date
    14 September 2026
  • 11.00 AM Eastern Time (US/Canada)
    03.00 PM GMT

Course is now LIVE. Click below to join the session.

This course strengthens the ability to assess product changes across connected technical documentation, helping teams identify where design modifications affect risk, clinical conclusions, verification evidence, and the regulatory record before inconsistencies become embedded.

/* Hide default + / - icon */ .accordion-toggle-icon { opacity: 0; } /* Add circle instead */ .accordion-header::after { content: "○"; font-size: 18px; margin-left: auto; transition: 0.2s ease; } /* When open → filled circle */ .accordion-item.active .accordion-header::after { content: "●"; } /* Hide default + / - icon */ .accordion-toggle-icon { opacity: 0; } /* Add circle instead */ .accordion-header::after { content: "○"; font-size: 18px; margin-left: auto; transition: 0.2s ease; } /* When open → filled circle */ .accordion-item.active .accordion-header::after { content: "●"; } .accordion-header::after { content: "◯"; font-size: 18px; transition: transform 0.25s ease; } .accordion-item.active .accordion-header::after { transform: rotate(90deg); }
REGISTRATION OPTIONS

Live session Plus
Complimentary 30 Days Streaming access

$290  |  One participant (viewer)

$390  |  Team (2 - 5 participants)

$490 |  Team (6 - 10 participants)

REGISTER FOR THE COURSE

US $290 per learner

30-Days Unlimited Streaming Access

/* Hide default + / - icon */ .accordion-toggle-icon { opacity: 0; } /* Add circle instead */ .accordion-header::after { content: "○"; font-size: 18px; margin-left: auto; transition: 0.2s ease; } /* When open → filled circle */ .accordion-item.active .accordion-header::after { content: "●"; } /* Hide default + / - icon */ .accordion-toggle-icon { opacity: 0; } /* Add circle instead */ .accordion-header::after { content: "○"; font-size: 18px; margin-left: auto; transition: 0.2s ease; } /* When open → filled circle */ .accordion-item.active .accordion-header::after { content: "●"; } .accordion-header::after { content: "◯"; font-size: 18px; transition: transform 0.25s ease; } .accordion-item.active .accordion-header::after { transform: rotate(90deg); }

REGISTER FOR THE COURSE

To Get 30-Day Access to ONLY this Course 

US $290 per learner
  • This course is Included in Subscription Pack
Subscription include access to entire Learning Library
/* Hide default + / - icon */ .accordion-toggle-icon { opacity: 0; } /* Add circle instead */ .accordion-header::after { content: "○"; font-size: 18px; margin-left: auto; transition: 0.2s ease; } /* When open → filled circle */ .accordion-item.active .accordion-header::after { content: "●"; } /* Hide default + / - icon */ .accordion-toggle-icon { opacity: 0; } /* Add circle instead */ .accordion-header::after { content: "○"; font-size: 18px; margin-left: auto; transition: 0.2s ease; } /* When open → filled circle */ .accordion-item.active .accordion-header::after { content: "●"; } .accordion-header::after { content: "◯"; font-size: 18px; transition: transform 0.25s ease; } .accordion-item.active .accordion-header::after { transform: rotate(90deg); }
  • Faculty
    José Ignacio Mora
  • Duration
    90 Minutes
  • Course ID
    TF1385
  • Live Q&A +
    Post-live Continued Learning
  • Presentation Handout
    & Templates
  • Assessment
    & Certification Included


Why take this course?

EU MDR technical documentation must remain internally consistent as a medical device changes through its lifecycle. A modification to design, materials, software, manufacturing, suppliers, intended use, or performance can affect risk controls, verification and validation evidence, clinical conclusions, labeling, and other records that support the approved device configuration. When these elements are maintained separately, individually controlled documents can still describe different product states.


A configuration-centered approach treats technical documentation as connected information rather than isolated files. This course focuses on identifying relationships among design information, risk controls, clinical evidence, verification evidence, and approved changes so that impact can be assessed before records drift apart. It addresses how applicability, effectivity, revision state, and objective evidence help determine whether existing conclusions remain valid after change, while supporting coherent technical documentation for EU MDR conformity assessment, Notified Body review, and ongoing regulatory oversight.

Follow Change Impact Across the Technical Documentation 

Product changes can create regulatory exposure when affected records are identified by document name rather than by their relationship to the changed product configuration. Develop a more disciplined way to follow change impact across design information, risk controls, clinical evidence, verification and validation, labeling, manufacturing information, and supporting regulatory records

Determine Whether Existing Evidence Still Applies

A current document does not guarantee that its evidence applies to the current device configuration. Strengthen the ability to question revision state, applicability, effectivity, and supporting evidence so that risk conclusions, clinical support, verification results, and design records remain aligned when technical documentation is reviewed.

Key Areas Covered

  • Relationships among design requirements, risk controls, clinical evidence, verification and validation records
  • Product configuration as the basis for maintaining consistent EU MDR technical documentation
  • Change impact across materials, software, suppliers, manufacturing processes, design characteristics, intended use, and performance
  • Applicability and effectivity of existing evidence after an approved product or process change
  • Identifying missing, stale, conflicting, or configuration-wrong relationships across controlled information
  • Alignment of risk conclusions, clinical support, design records, labeling, and verification evidence
  • Configuration-centered change assessment before approval rather than document-by-document reconstruction
  • Technical documentation consistency for EU MDR conformity assessment, Notified Body review, and regulatory oversight

Who Must Attend

  • Regulatory Affairs professionals
  • Quality Assurance and Quality Engineering
  • Design Assurance and Design Engineering
  • Risk Management professionals
  • Clinical Affairs and Clinical Evaluation personnel
  • Verification and Validation professionals
  • Systems and Software Engineering
  • Manufacturing and Process Engineering
  • Configuration and Change Management
  • Document and Information Management
  • Compliance and Internal Audit professionals
  • Managers responsible for EU MDR technical documentation and product change control
Meredith Crabtree
COURSE DIRECTOR

José Mora

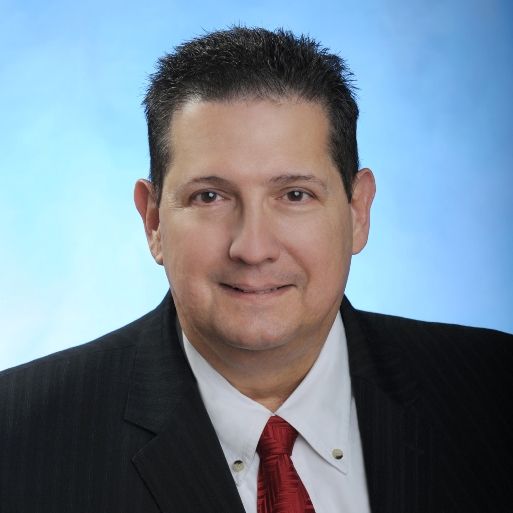
José Ignacio Mora brings more than 30 years of medical device and life sciences experience spanning quality systems, manufacturing engineering, design engineering, process development, validation, configuration control, and controlled-document methodology. His background includes leadership roles at Boston Scientific, Stryker Orthopedics, UroSurge, and Cordis, with direct experience in product development, manufacturing change, qualification, and quality-system improvement.

If you would like to request a Proforma invoice to sign up for this course. please click here

Commonly Asked Questions About This Subject

The following questions address practical regulatory, compliance, validation, quality, operational, and inspection-related considerations commonly associated with this subject.

How can technical documentation be current and still fail to represent the current device correctly?

A document can be current, approved, and properly controlled while the technical documentation as a whole remains inconsistent. Reviewers are concerned with whether the evidence supports the device configuration being presented. If a design output changed but the risk control, verification result, clinical conclusion, or labeling still reflects an earlier state, document status alone does little to resolve the inconsistency.


A useful review follows the relationships created by the change. Which requirement or design output changed? Which risks or controls depend on it? What evidence demonstrated that control or characteristic? Does that evidence still apply to the revised configuration? Are clinical conclusions or claims dependent on the changed characteristic?


This is where revision state, applicability, effectivity, and stable relationships become important. Technical documentation is easier to defend when the organization can show why each relevant item of evidence belongs to the configuration under review, rather than simply showing that every individual file passed document control.

How far should a change impact assessment extend when the original design change appears relatively minor? 

Start with the characteristic that changed and follow its dependencies before deciding which files need revision. A material change, software modification, supplier change, manufacturing change, or revised design requirement can affect several controlled relationships even when the initial change request appears narrow.


The assessment should identify the applicable requirement or design output, associated risks and risk controls, verification or validation evidence, and any clinical or performance conclusions that depend on that product characteristic. It should also establish which product revisions or configurations are affected. That prevents evidence from an earlier configuration from being carried forward merely because the corresponding report remains approved.


A weak assessment often becomes visible later as conflicting revisions, unsupported conclusions, or evidence that cannot be clearly tied to the changed device. Following the affected configuration objects and relationships before approval makes the impact visible while there is still an opportunity to address it systematically.

When can existing risk, verification, validation, or clinical evidence continue to support a changed device?

Existing evidence can remain usable when the organization can show that the change does not invalidate the conditions, characteristics, or configuration for which that evidence was generated. The justification needs to connect the changed product information to the evidence rather than relying on the age or approval status of the report.


For verification or validation evidence, examine whether the tested configuration, method, acceptance basis, and affected design characteristics remain applicable. For risk information, determine whether the change alters a hazard, risk control, implementation of that control, or residual-risk conclusion. Clinical evidence requires similar attention when a changed characteristic affects the device features, intended purpose, performance, or claims supported by that evidence.


The difficult cases are those where a report looks relevant but was produced against a different revision or configuration. Applicability should be demonstrable from controlled relationships and configuration state, not reconstructed later from assumptions.

Why can a complete traceability matrix still be insufficient during review of a changed device? 

A traceability matrix can help a reviewer see relationships, but its completeness does not prove that the underlying relationships are current. A matrix is a view of information at a particular state. If a requirement, risk control, design output, verification method, or evidence package changes without the underlying relationship being maintained, the matrix can appear complete while representing the wrong configuration.


For change control, traceability becomes useful when it exposes what else may be affected before approval. A changed requirement should lead reviewers to the relevant risk controls, implementing design outputs, verification evidence, clinical implications where applicable, and the product configurations for which those relationships remain valid.


This matters during technical documentation review because a reviewer needs more than a chain of references. The references must point to the correct revisions and applicable evidence. Maintaining the underlying configuration relationships makes traceability a reliable output rather than a manually reconstructed record.

Your TalkFDA Webinar Experience

When you reserve your seat, everything is organized for you — from access to certification from one place.

1. Confirmation

You receive a confirmation email and your course appears instantly in your TalkFDA dashboard.

2. Your Course Hub
Your TalkFDA course page becomes your central hub where you can join the session, access materials, and manage your learning.

3. Join the Live Training
At the scheduled time, click Join Session and you’ll be connected to the live Webex session with the instructor.

4. Watch Again Anytime
After the session, the playback becomes available on the same course page so you can revisit important sections.
5. Earn Your Certificate
Complete the course and your certificate is unlocked automatically in your learning history.

Everything related to your webinar: access, materials, playback, and certification — lives in one place.
Simple. Organized. Professional.
Go ahead and reserve your seat with confidence.

Built on 20+ years of global regulatory training experience across FDA, EMA, MHRA, and ICH frameworks.
Trusted by professionals across 80+ countries.

Testimonials

This is an upcoming session. Feedback below reflects experiences from similar programs delivered by our expert faculty.

“Session was easy to follow even for non-core team members. That helped.”
- Production Officer
“Good balance. Not too basic, not too deep. Worked well for mixed team.”
- Manager, Regulatory Affairs
“Team found it useful. Especially for aligning understanding across functions.”
- Director, Operations

Ready to Strengthen Your Team? Let’s Build Your Training Plan.

Whether you’re looking for a single onsite workshop or a multi-team training series, we’ll help you design a program that fits your goals, timelines, and operational reality.

Your team deserves the clarity.
Your organization deserves the confidence.

Upcoming Courses

Featured Courses